Daniele Padula
Associate Professor · Dipartimento di Biotecnologie, Chimica e Farmacia, Università di Siena
I'm a computational chemist working on the electronic structure and excited-state dynamics of organic materials — charge and energy transfer in conjugated polymers and organic semiconductors, and structure-property relationships for organic photovoltaics. The Research page covers current themes in more detail (auto-generated from my publications).
Contact
Dipartimento di Biotecnologie, Chimica e Farmacia, Università di Siena
Via Aldo Moro 2, 53100 Siena, Italy
Recent highlighted publications
Auto-selected from the last 5 years: highest impact-factor and most-cited papers.
This section is generated automatically and not reviewed by hand. It may contain errors — let me know if you spot one.
Circularly Polarized Luminescence from an Inverted Singlet–Triplet Chiral Dye
Molecules with an inverted singlet–triplet gap (ΔEST= ES1– ET1< 0) hold potential for optoelectronic applications as OLEDs and photocatalysis. Despite growing interest, no single-molecule emission from a chiral dye with an inverted gap has been reported, and only one case has shown such emission from supramolecular aggregates. Here, we present the first circularly polarized light emission (CPL) from a chiral molecule exhibiting an inverted singlet–triplet gap. Our design is based on the achiral heptazine core, functionalized with chiral substituents. Computational screening confirmed that the inversion of the singlet–triplet gap is retained upon derivatization. Photophysical characterization, including temperature-dependent optical spectra and photoluminescence decay profiles, supported by computational results, confirms the inverted gap in both achiral and chiral derivatives. Chiroptical properties show dissymmetry factor glumvalues (≈ 10–3) comparable to other chromophores functionalized with chiral groups. Combined with promising photoluminescence quantum yields, these findings highlight the potential of such materials for circularly polarized OLED devices.
Toward Circularly Polarized Luminescence from Inherently Chiral Inverted Singlet–Triplet Chromophores
Recent years have seen rising interest in molecules that violate Hund’s rule, where the first excited singlet state lies below the triplet (ΔEST< 0), accelerating reverse intersystem crossing, a key process in thermally activated delayed fluorescence. We present a computational study on such inverted singlet–triplet (IST) molecules modified for chirality to enable Circularly Polarized Luminescence (CPL), an unexplored direction for IST emitters. Two main chromophores, triangulene and pentalene, were functionalized with minimal chiral groups, preserving negative ΔEST, as demonstrated via multiconfigurational calculations. We also explored inherently chiral, nonplanar chromophores, focusing on extended triangulenes resembling helicenes. Substituents enabling enantiomer separation were introduced, and racemization barriers were assessed. When singlet–triplet inversion was lost, structures were further optimized, yielding a promising substrate combining a high racemization barrier, strong CPL, and inverted singlet–triplet energetics, making it suitable for CP-OLED applications.
Identification of Unknown Inverted Singlet–Triplet Cores by High-Throughput Virtual Screening
Molecules where the energy of the lowest excited singlet state is found below the energy of the lowest triplet state (inverted singlet-triplet molecules) are extremely rare. It is particularly challenging to discover new ones through virtual screening because the required wavefunction-based methods are expensive and unsuitable for high-throughput calculations. Here, we devised a virtual screening approach where the molecules to be considered with advanced methods are pre-selected with increasingly more sophisticated filters that include the evaluation of the HOMO-LUMO exchange integral and approximate CASSCF calculations. A final set of 7 candidates (0.05% of the initial 15 000) were verified to possess inversion between singlet and triplet states with state-of-the-art multireference methods (MS-CASPT2). One of them is deemed of particular interest because it is unrelated to other proposals made in the literature.
Assessing alkyl side chain effects on electron transport properties of Y6-derived non-fullerene acceptors
The effect of alkyl side chains on the electron transport properties of non-fullerene acceptors is assessed, concluding that the main influence is on crystal packing rather than on dynamic disorder.
Organic materials repurposing, a data set for theoretical predictions of new applications for existing compounds
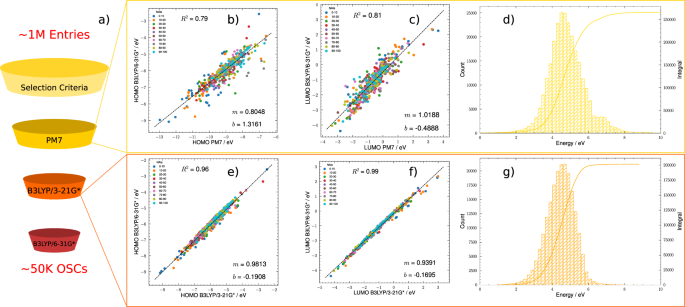
We present a data set of 48182 organic semiconductors, constituted of molecules that were prepared with a documented synthetic pathway and are stable in solid state. We based our search on the Cambridge Structural Database, from which we selected semiconductors with a computational funnel procedure. For each entry we provide a set of electronic properties relevant for organic materials research, and the electronic wavefunction for further calculations and/or analyses. This data set has low bias because it was not built from a set of materials designed for organic electronics, and thus it provides an excellent starting point in the search of new applications for known materials, with a great potential for novel physical insight. The data set contains molecules used as benchmarks in many fields of organic materials research, allowing to test the reliability of computational screenings for the desired application, “rediscovering” well-known molecules. This is demonstrated by a series of different applications in the field of organic materials, confirming the potential for the repurposing of known organic molecules.
Multiple charge separation pathways in new-generation non-fullerene acceptors: a computational study
Through a combination of molecular dynamics, density functional theory and Fermi's golden rule, computational modelling of two all-small-molecule organic bulk heterojunctions provides a simple rationalisation of the great performance of Y6.